Longevity Weekly Review 2026-10-14
Week In Review
This was a week when the clinic, not the mouse room, set the agenda. Three separate human efforts to rejuvenate aged tissue reported early data, and together they mark how quickly ideas that were preclinical a few years ago are now being tested in people. Rubedo Life Sciences showed that a topical senotherapeutic could measurably lower the epigenetic age of skin in a controlled Phase 1b trial. Life Biosciences reported the first in-human data from a gene therapy that switches on three of the Yamanaka reprogramming factors inside the eye, with no serious safety signals and early hints of restored vision. And at the heterodox end of the spectrum, the KHL Foundation described what it believes are the first administrations of the same reprogramming factors to living people, outside the trial system entirely. Read together in Rubedo Reports Younger Skin Age in a Phase 1b Trial, Partial Reprogramming Reaches a Human Trial in the Eye, and The First Human Partial Reprogramming, Outside the Trial System, these three stories bracket the range of how rejuvenation biology is actually entering human bodies, from the carefully regulated to the impatient.
The commercial and strategic framing of the field also shifted. A senior Novartis scientist made the case that drug developers should aim to regenerate aged tissue rather than merely slow its decline, arguing in The Case for Regenerating Tissue Rather Than Slowing Decline that restoration is easier to measure, easier to price, and more likely to win a regulatory path than incremental deceleration. That argument is echoed in the lab, where two mechanistic papers this week were about restoring lost capacity rather than preventing damage: a study linking the under-studied gene ZNF865 to the collapse of spinal discs in A Single Gene May Hold Spinal Discs Back From Aging, and a dissection of why the heart fails to repair itself in An Enzyme Switch That Governs Whether the Heart Can Repair Itself.
Senescent cells, the field’s most familiar target, were reexamined rather than simply attacked. One study found that cells made senescent by different triggers broadcast very different signals, complicating the idea of a single clearance drug, as described in Not All Senescent Cells Speak the Same Language. Another company, Immorta Bio, proposed teaching the immune system itself to recognize and remove those cells with a synthetic peptide designed by artificial intelligence, covered in An AI-Designed Peptide That Trains the Immune System to Clear Aged Cells.
Finally, two careful metabolic studies refined long-held assumptions. One argued that the celebrated lifespan benefits of methionine restriction are really about a different amino acid, cysteine, as laid out in The Longevity Diet May Have Been About the Wrong Amino Acid. The other traced how a single molecule made by a gut bacterium acts like a natural statin to slow arterial disease, reported in A Gut Bacterium That Makes Its Own Statin. Across all ten items, the common thread is specificity: the field is moving from blunt interventions toward named targets, named cell populations, and measurable endpoints in actual people.
Items
Rubedo Reports Younger Skin Age in a Phase 1b Trial

Most interventions that claim to reduce biological age are measured in cells or mice. Rubedo Life Sciences this week reported that its topical drug RLS-1496 did so in people, in a controlled Phase 1b study. According to Longevity.Technology, four weeks of treatment in patients with atopic dermatitis lowered the measured epigenetic age of sun-damaged skin by an average of 2.1 years at the follow-up point, with the best responder showing a reduction of nearly twelve years.
The drug is what Rubedo calls a “SenoAdaptive” senotherapeutic. Rather than killing every stressed cell, it selectively pushes genuinely pathological senescent cells into a form of iron-dependent cell death called ferroptosis by targeting the enzyme GPX4, while nudging merely stressed but still useful neighboring cells to switch on their own antioxidant defenses. The intent is to remove the harmful cells without the collateral damage that cruder senolytics risk.
The company reported supporting signals beyond the aging clock. In a separate study of actinic keratosis, a precancerous skin condition, 22 patients saw an average 49 percent reduction in lesions after 28 days. Blood analysis of roughly 10,800 proteins showed a 17 percent drop in an inflammatory marker tied to atopic dermatitis in treated patients, against a 19 percent rise in those given placebo, and skin biopsies showed reduced senescence signatures alongside a 10 percent increase in type III collagen.
These are small, early-stage numbers, and skin is an unusually accessible tissue that may flatter any topical approach. But the result matters because it couples a mechanistic story about which cells are removed with a clinical readout in humans. Chief executive Frederick Beddingfield framed the data as evidence that the approach could reach “beyond dermatology,” and the company has a Phase 2a study in hair loss planned next.
Source: Longevity.Technology
Partial Reprogramming Reaches a Human Trial in the Eye

Cellular reprogramming, the idea of winding back a cell’s apparent age by briefly expressing the genes that make embryonic stem cells, has been one of longevity biology’s most exciting and most theoretical ideas. This week Life Biosciences reported the first human data from a trial that puts it to the test in a real disease. Its therapy, ER-100, delivers three of the four Yamanaka factors, OCT4, SOX2, and KLF4, into the eye by injection, with the aim of restoring function to the retinal ganglion cells damaged in glaucoma.
The company reported that three participants with open-angle glaucoma received the starting dose of 2×10¹¹ vector genomes. Through 56 days, the treatment was well tolerated, with no dose-limiting toxicities, no serious adverse events, and no safety signals attributed to the therapy itself. Expression of the reprogramming factors is held under control by an oral drug, doxycycline, so the rejuvenation program can be switched off, a safety feature given the theoretical risk that reprogramming taken too far could produce tumors.
More striking than the safety data was a hint of benefit. Two of the three participants showed improvement in formal visual-field testing, with at least 28 percent of tested points improving by more than four decibels. An independent safety board reviewed the results and cleared the trial to escalate to a threefold higher dose.
Three patients and two months of follow-up are a signal, not a proof. But the significance is conceptual as much as clinical: this is partial reprogramming leaving the mouse colony and entering a regulated human trial, with the fine-grained safety controls that such a powerful tool demands. Chief Scientific Officer Sharon Rosenzweig-Lipson described the data as early information on tolerability and on epigenetic restoration in people, deliberately modest language for a field prone to overreach.
Source: Life Biosciences
The First Human Partial Reprogramming, Outside the Trial System
The same reprogramming factors reached human beings this year by a very different route, and the contrast is instructive. The KHL Foundation, a small organization founded by two people in their eighties, reported that it had administered an experimental eleven-gene therapy carrying the OCT4, SOX2, and KLF4 factors to a human recipient in March 2026, outside the conventional clinical-trial system. The longevity writer at Fight Aging treats the episode as a landmark worth watching, while being careful to separate the milestone from any claim of proven benefit.
The foundation’s own account describes a deliberate philosophy it calls “Develop, Apply, Monitor, Adapt,” aimed at people who feel they do not have decades to wait for the formal development pipeline. Alongside the reprogramming therapy, its founders reported taking a separate three-gene “rejuvenation cocktail” of follistatin, klotho, and SIRT1, and described anecdotal changes such as increased strength and a fall in fasting blood glucose. They stress these are observations from the first two recipients, not controlled results, and do not establish cause.
The scientific caution here cannot be overstated. There is no control group, no peer review, and no way yet to know whether the therapy did anything at all, let alone whether it was safe over the long term. Medical-tourism longevity treatments have a poor record of generating usable data.
The item earns attention precisely because it sits at the field’s frontier of impatience. It raises, in concrete form, the questions that the regulated trials will spend years answering carefully: how to express reprogramming factors safely in a living adult, how to measure whether anything was rejuvenated, and how a field should respond when people decide not to wait. The responsible path runs through trials like Life Biosciences’, but the pressure that produces efforts like this one is real and worth understanding.
Source: Fight Aging
An AI-Designed Peptide That Trains the Immune System to Clear Aged Cells
Most attempts to remove senescent cells use drugs that kill them directly. Immorta Bio this week described a different strategy: rather than delivering the killer, teach the body’s own immune cells to do the job. Its candidate, called SenoMune-0257, is a short synthetic peptide, just nine amino acids long, designed to stimulate cytotoxic T cells to recognize and eliminate senescent cells while leaving healthy ones alone.
The approach is a bet that the immune system can be pointed at aged cells the way it is increasingly pointed at cancer. As people age, immune surveillance of senescent cells declines, allowing them to accumulate and spread inflammatory signals. A peptide that restores that recognition, and leaves behind durable immunological memory, could in principle keep clearing senescent cells long after the drug itself is gone, something no conventional senolytic can claim.
What makes the candidate notable is how it was found. Immorta Bio reports that the peptide was designed with artificial intelligence, and that building it as an off-the-shelf synthetic molecule rather than a personalized cell therapy gives it large advantages in manufacturing cost and scalability. The company says it has completed preclinical studies, with academic collaborators including the Buck Institute, Cedars-Sinai, UC San Diego, and the University of Miami, and plans to move toward clinical development.
The candidate remains investigational, with no established human safety or efficacy, and the leap from a designed peptide to a working immunotherapy is large. But the item illustrates two converging trends worth tracking: the use of machine learning to design biological molecules, and the shift from killing senescent cells with chemistry to recruiting the immune system against them.
Source: Longevity.Technology
The Case for Regenerating Tissue Rather Than Slowing Decline

Much of the longevity field frames its goal as slowing the rate of decline. Fiona Marshall, who leads biomedical research at Novartis, made the case this week that large drug developers should instead aim to regenerate aged tissue, and her reasoning is as much commercial and regulatory as scientific. Her argument matters because it comes from inside one of the world’s largest pharmaceutical companies rather than a longevity startup.
Marshall’s first point is a counterintuitive one about measurement and cost. A drug that restores lost function, she argues, may actually be cheaper to develop than one that slows a decline, because “it’s easier to measure an improvement than a reduction in a decline.” A trial that can show a tissue getting better needs fewer patients and less time than one trying to prove that something got worse more slowly, which bears directly on whether such drugs ever reach patients.
She also pushes against one-size-fits-all thinking. Novartis’ aging and regenerative medicine division organizes its work around three underlying processes, mitochondrial dysfunction, declining DNA repair, and changes in the extracellular matrix that holds tissues together. And she argues for identifying “aging endotypes,” distinct biological pathways by which different people age, so that someone with inflammation-driven aging receives a different therapy than someone whose aging is metabolically driven.
Marshall is candid that regulation is the gating factor, saying that a clear FDA roadmap for developing an antiaging drug would bring the industry “right in there.” The toolkit she describes spans small molecules, RNA-based drugs, gene therapy, and partial cellular reprogramming. The significance of the piece is that it reframes longevity as a tractable drug-development problem with ordinary commercial logic, not a moonshot, and does so from a position of considerable industrial weight.
Source: Longevity.Technology
A Single Gene May Hold Spinal Discs Back From Aging
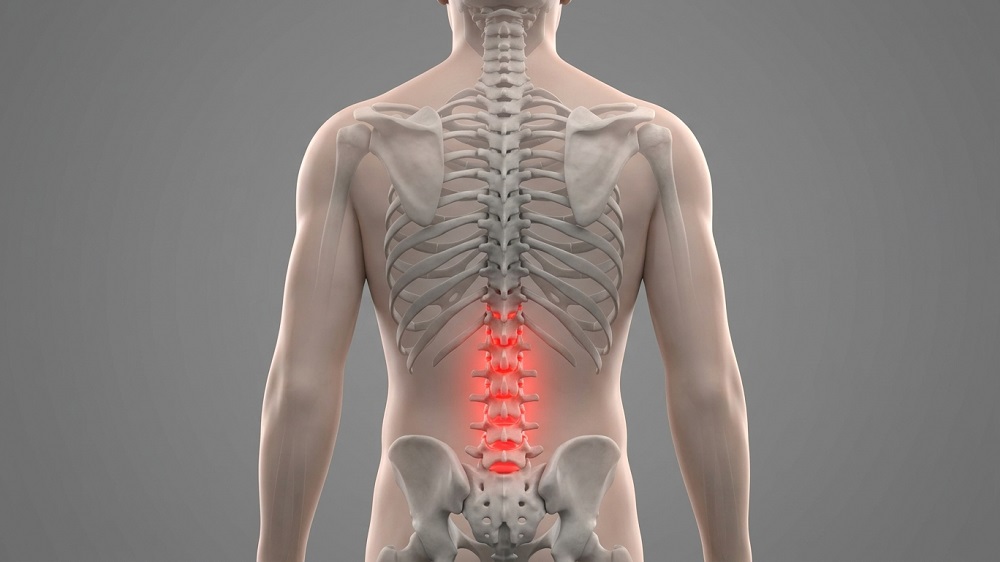
Degeneration of the spinal discs is one of the most common and least glamorous features of aging, a leading cause of back pain and disability. Researchers reporting in the journal Aging Cell this week identified a little-studied gene, ZNF865, as a surprisingly central regulator of whether the cells inside those discs stay healthy or slide into senescence.
The team found that disc cells from degenerated human tissue carried much lower levels of ZNF865 than healthy cells. When they silenced the gene in healthy cells, those cells showed roughly five times the markers of senescence, along with the inflammatory secretions that drive tissue breakdown. Running the experiment in reverse, boosting ZNF865 in already-degenerated cells pushed their senescence markers back down toward healthy levels and restored their production of collagen, the structural protein discs need.
The link held up in animals. Rats that received disc cells depleted of ZNF865 developed the hallmarks of degenerative disc disease within about two months, suggesting the gene’s loss is not just a marker of damage but a driver of it.
The researchers are candid that the detailed molecular mechanism remains unknown, and that no drug targeting ZNF865 exists; so far the gene can only be manipulated with gene-editing tools in the lab. But identifying a single control point that sits upstream of both senescence and the loss of structural repair gives the field a concrete new target in a tissue where the main current option is surgery.
Source: Lifespan.io
A Gut Bacterium That Makes Its Own Statin
Statins are among the most widely prescribed drugs in the world, and this week researchers reporting in Nature described a gut bacterium that appears to make a natural version of one. The bacterium, Bacteroides uniformis, is less abundant in people with atherosclerotic cardiovascular disease, and the team traced its benefit to a specific molecule it produces: pentadecanoic acid, a 15-carbon saturated fatty acid known as C15:0.
In mice bred to be prone to atherosclerosis, a twelve-week course of the bacterium reduced the area of arterial plaque, lowered LDL and total cholesterol, and brought down triglycerides. The researchers established that these effects were due to the C15:0 molecule itself, which works through the same biochemical lever as pharmaceutical statins: it inhibits HMG-CoA reductase, the enzyme that controls how much cholesterol the liver makes, and in turn increases the liver’s uptake of LDL from the blood.
The magnitude was meaningful if not dramatic. The molecule reduced plaque burden by about 50 percent in one experiment, though a conventional statin, atorvastatin, performed noticeably better at comparable dosing. Importantly, the effect was a slowing of plaque progression rather than a reversal of existing disease.
The work is still entirely in mice, and a dietary molecule is unlikely to replace a potent drug. But it is a clean example of the kind of mechanistic insight the field increasingly values: a named microbe, a named molecule, and a well-understood pathway connecting the gut microbiome to one of the central diseases of aging. It also hints at why the composition of the microbiome, which shifts with age, may influence cardiovascular risk in the first place.
Source: Lifespan.io
The Longevity Diet May Have Been About the Wrong Amino Acid
For years, restricting the amino acid methionine has been one of the most reliable ways to extend lifespan in laboratory animals, and a staple of discussions about dietary longevity. A study highlighted this week argues that the credit has gone to the wrong molecule. The real driver, the researchers contend, is restriction of a different amino acid, cysteine.
The distinction is easy to miss because the two are linked. The body makes cysteine from methionine, so a low-methionine diet is also, downstream, a low-cysteine diet. Standard methionine-restriction experiments, the authors note, remove both amino acids at once, confounding the interpretation. By separating the two, the team found that selectively cutting cysteine extended lifespan and improved stress resistance in worms independently of methionine levels, and that in mice engineered to lack a key enzyme, the metabolic benefits tracked cysteine rather than methionine.
In aged mice, a roughly 70 percent reduction in cysteine reprogrammed what the authors call the immunometabolic axis, the interplay between metabolism and the immune system, and improved measures of healthspan. The finding reframes a well-worn intervention around a more precise target.
That precision matters for anyone hoping to translate dietary findings into a therapy. Knowing which molecule actually carries the effect is the difference between a vague recommendation to eat less of certain proteins and a defined target that a drug or a carefully designed diet could act on. It is a reminder that even the field’s established results can rest on assumptions worth re-testing.
Source: Fight Aging
An Enzyme Switch That Governs Whether the Heart Can Repair Itself
The adult human heart barely regenerates, which is why a heart attack leaves permanent scarring instead of new muscle. Researchers reporting in Nature Cardiovascular Research this week identified a metabolic switch, centered on the enzyme succinate dehydrogenase, that helps determine whether damaged heart tissue repairs itself or stiffens into fibrosis.
Using single-cell sequencing to read the activity of individual cell types in injured hearts, the team found that inhibiting succinate dehydrogenase, either with a compound called malonate or by deleting the gene, shifted cells toward a more regenerative metabolic state and remodeled the way their DNA was packaged. The intriguing result was that the benefit depended entirely on which cells were affected. Switching the enzyme off in the heart’s muscle cells made them divide more but did not improve function after a simulated heart attack. Switching it off in myofibroblasts, the cells that lay down scar tissue, suppressed fibrosis and improved the heart’s performance.
The lesson is that regenerating an organ is not simply a matter of making its primary cells multiply. Functional recovery required coordinating changes across several cell types at once, with the suppression of scarring mattering as much as the proliferation of muscle.
That insight cuts against a tempting simplification in regenerative medicine, the idea that coaxing the main working cells to divide will restore a damaged organ. By pinpointing a single enzyme whose effects depend on cellular context, the study offers both a candidate target and a cautionary principle: in the aging heart, the cells that build scars may be the more important ones to reprogram.
Source: Fight Aging
Not All Senescent Cells Speak the Same Language
Senescent cells are blamed for much of the inflammation that accompanies aging, and clearing them is one of the field’s main strategies. A study published in the journal Aging this week complicates that picture in a useful way, showing that cells pushed into senescence by different triggers broadcast strikingly different signals.
The researchers compared human fibroblasts made senescent by an activated cancer gene, RAS, against cells marked by the common senescence protein p16. The RAS-driven cells behaved as expected, pumping out inflammatory molecules and numerous small membrane-bound packets called extracellular vesicles. The p16 cells, by contrast, did not secrete the classic inflammatory cytokines at all, yet still released vesicles. Those vesicles were far from inert: when taken up by healthy young cells, they induced senescence in the recipients and caused DNA damage, and vesicles from the cells of older people drove young cells into senescence.
The authors propose splitting the familiar “senescence-associated secretory phenotype” into two parts, a soluble fraction of proteins and a separate vesicle fraction, because the two can be present independently and may do harm through different means. The vesicle cargo appears to be genotoxic, though its exact contents are not yet identified.
For the project of removing senescent cells, this is a clarifying warning. If different senescent cells signal differently, and some spread aging through vesicles without the inflammatory signature that most detection methods look for, then a single drug aimed at one senescent phenotype may miss others entirely. The path forward likely runs through matching the intervention to the specific kind of senescent cell, rather than treating them as a single enemy.
Source: Lifespan.io